Research Articles

Accurate Exciton Binding Energies in Organic Crystals: A Comprehensive Guide to GW-BSE Calculations for Biomedical Materials
This article provides a detailed exploration of the GW approximation and Bethe-Salpeter equation (GW-BSE) method for calculating exciton binding energies in organic molecular crystals.
GW-BSE Excited-State Geometry Optimization: A Practical Guide for Accurate Photochemical Predictions
This article provides a comprehensive, expert-level guide to performing and validating GW-BSE excited-state geometry optimizations for computational chemistry and drug discovery.
Benchmarking GW-BSE Excitation Energies: A Comparative Analysis of the Quest-3 Database for Molecular Photophysics
This article provides a comprehensive analysis of the GW-Bethe-Salpeter Equation (GW-BSE) approach for calculating molecular excitation energies, benchmarked against the extensive Quest-3 database.
Demystifying GW-BSE: Breaking the O(N⁶) Bottleneck for Accurate Drug Discovery
This article provides a comprehensive guide for researchers and drug development professionals on tackling the prohibitive O(N⁶) computational cost scaling of the GW approximation and Bethe-Salpeter Equation (BSE) method—a gold...
GW-BSE vs CASPT2 vs CC2: Benchmarking Excited State Energies for Drug Discovery
This article provides a comprehensive benchmark analysis of three advanced electronic structure methods—GW-BSE, CASPT2, and CC2—for calculating excited state energies.
Beyond the Band Gap Blind Spot: Correcting GW-BSE Underestimation for Accurate Pharmaceutical Material Discovery
This article addresses the critical challenge of band gap underestimation in the widely-used GW approximation and Bethe-Salpeter Equation (GW-BSE) method for computational materials science.
GW Starting Point Dependence: How to Choose the Right DFT Functional for Accurate Biomolecular Simulations
This article provides a comprehensive guide for researchers and drug development professionals on navigating the critical challenge of starting-point dependence in GW calculations.
GW Approximation Explained: Calculating Accurate Quasiparticle Energies for Materials & Drug Discovery
This article provides a comprehensive guide to the GW approximation for calculating quasiparticle energies in materials science and computational chemistry.
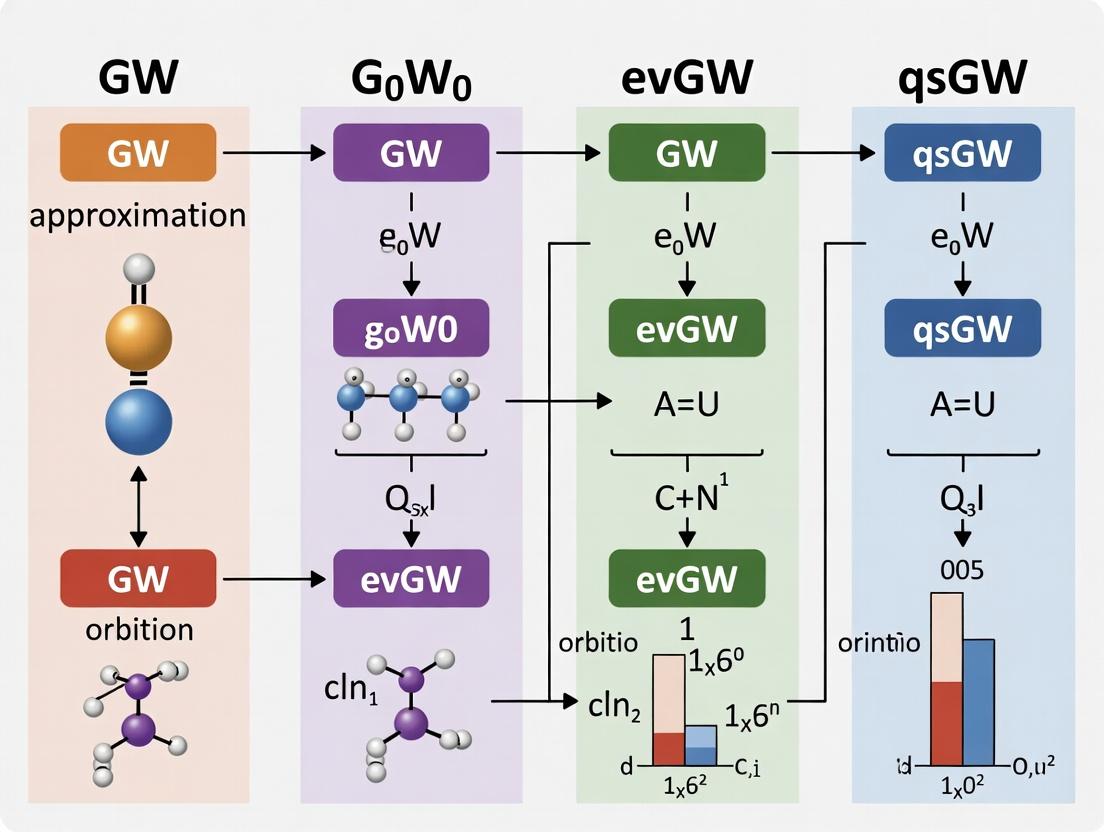
GW Approximation Decoded: A Comprehensive Guide to G0W0, evGW, qsGW, and Self-Consistency for Materials Science
This article provides a comprehensive overview of the GW approximation for quasiparticle energy calculations, with a focus on comparing the different levels of self-consistency: G0W0, evGW, qsGW, and scGW.
GW, G0W0, evGW, qsGW: A Comprehensive Benchmark for Accurate Ionization Potential Prediction in Molecular Systems
This article provides a detailed analysis and benchmark of four key GW approximation variants—G0W0, evGW, qsGW, and GW—for calculating molecular ionization potentials.